Computational chemistry, without the old workflow.
Run DFT with real scientific engines, and keep every result tied to its method, inputs and artifacts. An AI scientist plans and explains the work, but it cannot write a result.
Fig. 1Reference crystal structureCOD 9014036 · H. Sawada, Mater. Res. Bull. 29, 239–245 (1994)α-Cr₂O₃ (corundum type) · R-3c (No. 167) · a = b = 4.9570 Å, c = 13.5923 Å, γ = 120° · shown as a 2×2×1 display supercellIndependently sourced reference structure. It is not the refined structure of any ChemDFT sample or of any specific study.
Draw or import a structure. Each calculation points to the exact, immutable structure version it used.
Recorded executionWater · HF / STO-3G · PySCF 2.14.0
Structure
Water, H₂O
Atoms
3 · Cartesian, Å
Source
ChemDFT test geometry
Record
Immutable structure version
Input structure SHA-256
39e5a504cab0…55847b
State the chemistry before computing it
Charge, multiplicity, method and basis set are explicit. The normalised calculation input is hashed.
Recorded executionWater · HF / STO-3G · PySCF 2.14.0
Charge · multiplicity
0 · 1stated, not inferred
Method · basis
Restricted Hartree-Fock · STO-3G
SCF convergence
1×10⁻¹⁰ · max 100 cycles
A real engine does the calculation
This recorded attempt ran PySCF in an isolated worker. The engine produced the result; the browser and AI did not.
Recorded executionWater · HF / STO-3G · PySCF 2.14.0
Engine
PySCF 2.14.0
Execution
Isolated worker process
Attempt
1 · succeeded in 0.63 s
Started
2026-09-27 14:19:44 UTC
Queued
Running
Succeeded
Results are checked before they are trusted
The platform checks convergence, output validity and producer authority before storing the result. Scientific review remains separate.
Recorded executionWater · HF / STO-3G · PySCF 2.14.0
SCF
converged · 7 cycles
Total SCF energy
-74.9629282310 Eh
Electrons
10 · 5 α / 5 β
Basis functions
7 · 7 molecular orbitals
Status
Calculated · not reviewedcomputed does not mean checked by a person
Inspect what the calculation produced
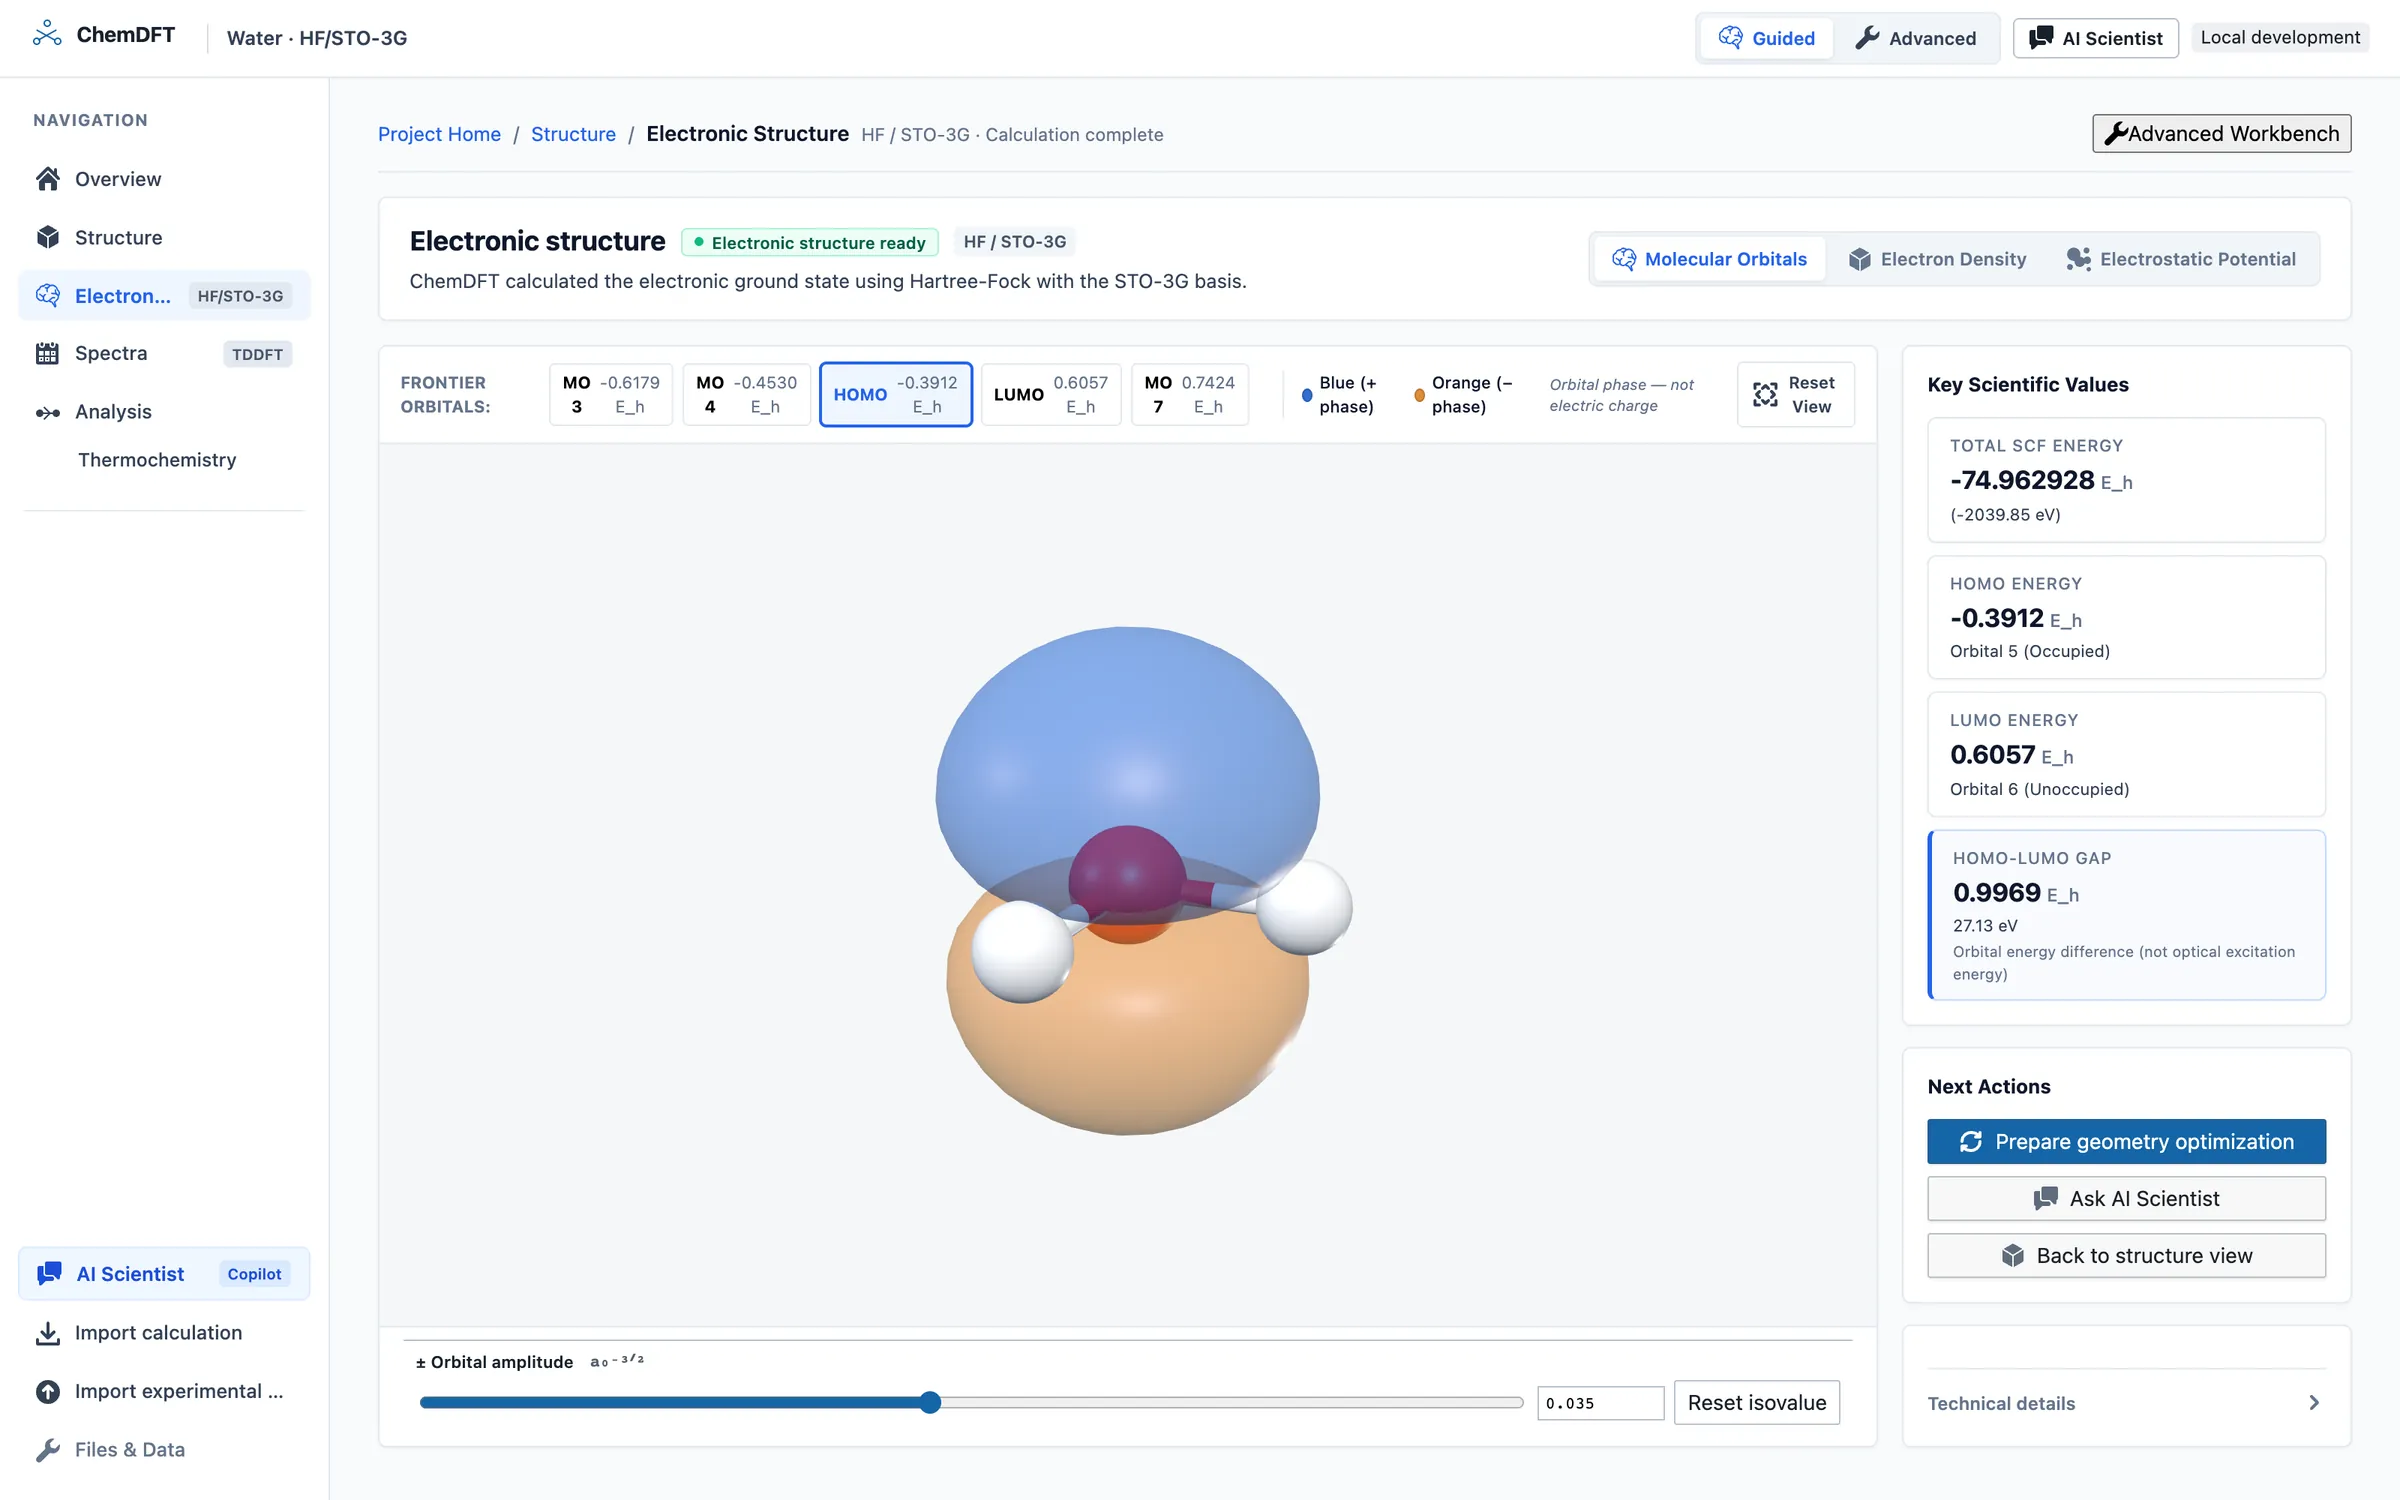
Inspect the stored water orbitals and their energies in the workspace, alongside the method and evidence that produced them.
Recorded executionWater · HF / STO-3G · PySCF 2.14.0
HOMO · MO 5
-0.39124467 Eh-10.64631 eV
LUMO · MO 6
0.60567399 Eh16.48123 eV
HOMO–LUMO gap
0.99691866 Eh27.12754 eV · orbital-energy difference; not an excited-state calculation.
Recorded executionWater · HF / STO-3G · PySCF 2.14.0
Structure
Water, H₂O
Atoms
3 · Cartesian, Å
Source
ChemDFT test geometry
Record
Immutable structure version
Input structure SHA-256
39e5a504cab0…55847b
A recorded execution, not a live calculation. Fields come from the same stored run throughout.
What runs your calculation
Engines
ChemDFT calls these projects; it does not reimplement them, and none of them endorse it. Not every engine supports every workflow, and limits below are those of the hosted worker.
Molecular calculations
PySCF
● Available
Quantum chemistry for molecules
Version 2.14.0
Licence: Apache-2.0
HF, B3LYP and PBE with STO-3G, 6-31G, def2-SVP and def2-SVPD, in the gas phase.
Energy, geometry optimisation, frequencies, orbitals, TDDFT and thermochemistry; Libxc and libcint supply functionals and integrals.
Hosted size limits are tighter: 12 atoms for optimisation and orbitals, 6 for frequencies and TDDFT.
geomeTRIC
● Available
Geometry optimisation
Version 1.1.1
Licence: BSD-3-Clause plus a no-AI-training clause (custom)
Optimises molecular geometries using PySCF gradients.
Periodic structures are not optimised with it.
xTB (GFN2)
● Available
Semi-empirical energy and optimisation
Version 6.7.1
Licence: LGPL-3.0-or-later
GFN2-xTB energy and native geometry optimisation for molecules.
Hosted limit: 50 atoms. Hosted runs so far are few, with no independent Linux reference.
CREST
● Available
Conformer search
Version 3.0.2
Licence: GPL-3.0 (with LGPL-3.0 components)
GFN2 conformer search for neutral H, C and O singlets; hosted limit 12 atoms.
The search is stochastic; retained conformers are not certified minima.
Periodic crystals and solids
Quantum ESPRESSO
● Available
Plane-wave periodic DFT
Version 7.6 · SSSP PBE Efficiency v2.0
Licence: GPL-2.0-or-later
PBE or PBE+U (Dudarev Hubbard U); non-magnetic, ferromagnetic or antiferromagnetic; k-point meshes from Γ up to 4×4×4.
Optional relaxation of atom positions with the cell held fixed, density of states and band structure.
Fixed pseudopotentials and cutoffs; at most 20 atoms. No cell relaxation, stress, phonons, hybrid functionals or spin–orbit coupling.
PySCF periodic
● Available
Gaussian-basis periodic DFT
Version 2.14.0
Licence: Apache-2.0
Single-point PBE or PBE+U energy with a Gaussian basis (GTH-PBE pseudopotentials); non-magnetic, ferromagnetic or antiferromagnetic.
Γ point only on the hosted worker; at most 20 atoms.
No relaxation, bands, density of states, forces or stress.
Structures and imports
RDKit
● Available
Molecular structure handling
Version 2025.9.1
Licence: BSD-3-Clause
Reads and represents molecular structures.
Does not run calculations.
pymatgen
● Available
Crystal structures from CIF
Version 2025.10.7
Licence: MIT
Reads crystal structures from CIF and builds the band-structure path for Quantum ESPRESSO.
Does not run calculations.
cclib
● Available
Parsing imported output
Licence: BSD-3-Clause
Parses imported Gaussian 09/16 and ORCA 5.x–6.1 output.
Not available yet
Psi4
○ Not available
Quantum chemistry
Licence: LGPL-3.0-only AND BSD-3-Clause AND MIT
Psi4 1.11 single-point energy for five accepted method/basis pairs (HF, B3LYP, PBE, MP2); neutral singlets only. Not available: there is no Linux build of Psi4 yet and some of its licence items are open decisions
DFT-D4
○ Not available
Dispersion correction
Licence: LGPL-3.0-or-later
DFT-D4 4.2.0 dispersion correction of accepted Psi4 B3LYP or PBE def2-SVP energies; combined totals are not independently verified. Not available: it accepts only a Psi4 energy, and none can be produced here
GPU4PySCF
○ Not available
GPU acceleration
Licence: Apache-2.0
GPU acceleration. Not available: no program it could drive has a hosted workload
QCElemental · QCEngine · QCFractal · Basis Set Exchange · ASE
○ Not available yet
Interchange and orchestration
Licence: Differs per project; see the licence table
Interchange and orchestration, not integrated
nmrglue · pyOpenMS
○ Not available yet
Spectroscopy and data
Licence: Licence not yet recorded
NMR and mass-spectrometry data, not integrated
Other limits
New Psi4 and DFT-D4 runs: Psi4 isn't configured on this deployment
Psi4 optimisation, frequencies or charged and open-shell systems
Periodic cell relaxation, stress, phonons, hybrid functionals and spin–orbit coupling
Molecular dynamics and protein-scale simulation
Every functional, basis set or quantum method
Gaussian input-file compatibility
Bring-your-own-key AI, public collaboration
Public sign-up and pricing
Also available
Structure import (XYZ, MOL, SDF, CIF) and 2D drawing
Orbital, vibrational, IR/Raman and thermochemistry views
Experimental spectrum import and comparison
An AI Scientist that works from stored results, with citations
ORCID sign-in for accounts that already linked an ORCID iD in Settings
Each release lists what was and was not verified, and on which platform, in the release notes. A stored run record is shown under Reproducibility. The one measured agreement check is on the Validation page. The open-core source snapshot for release 0.0.52 is on Zenodo, DOI 10.5281/zenodo.24209616 (all versions: DOI 10.5281/zenodo.24082321); it does not by itself verify any scientific result.
Worked example
One stored calculation: α-Cr2O3
CALCULATED ChemDFT 0.0.46 · Linux x86_64 · pilot EC2, 13 Nov 2026. Job 5682273d-6bcc-45c0-8dd1-ab2bf07b426a.
PBE+U, U_eff(Cr) = 3.7 eV (platform default, used_default_u = true)
Converged
yes; relaxation converged in 3 ionic steps, total force 0.001093 Ry/bohr
Final total energy
−475.99968609 Ha
Löwdin local Cr moments
±2.9995 μB (antiferromagnetic, total magnetization ≈ 0)
Kohn–Sham band gap
3.239 eV
Runtime
463.6 s
Same system, different settings (job 58a06173-0f87-46c1-b640-daa5c0803468): Γ point only, U_eff(Cr) = 2.0 eV; Kohn–Sham gap 2.896 eV, local Cr moments ±3.0028 μB.
The gap is a Kohn–Sham PBE+U eigenvalue gap, not an optical or experimental gap.
U is a platform default, not fitted.
This one calculation is not a benchmark.
Status CALCULATED with Quantum ESPRESSO 7.6 on this platform only (Linux x86_64, pilot EC2); it is not evidence for any other platform.
Licence
Licences
All of ChemDFT's own code is licensed under Apache-2.0: the web app, the API, the engine wrappers, the schemas and contracts, the infrastructure code and the documentation. Third-party engines and libraries keep their own licences.
ChemDFT calls 39 third-party engines and libraries and does not reimplement them; each stays under its own licence, and some licences are not yet recorded. See the full licence table
Scientific Result Firewall
AI may explain a scientific result. It cannot replace the calculation that produced it.
Authoritative values come from deterministic scientific engines and validated outputs, each stored with its calculation source: method, basis set, engine and engine version. The model works through the same typed actions as a person and has no way to write a result.
Platform · deterministic · the only path that writes a result
PlatformAuthoritative resultValidated, with provenanceWater · HF / STO-3G · PySCF 2.14.0 Stored · calculated · not reviewed4
AIAI interpretationExplains the stored result5
The AI Scientist can
Plan a workflow and choose which tool to propose
Explain a method, a basis set or a convergence message
Interpret a stored, validated result and cite it
Help troubleshoot a failed or unconverged calculation
The AI Scientist cannot
Write an energy, spectrum, orbital or any other authoritative value
Stand in for a calculation that did not run
Present an estimate as if an engine produced it
Change a stored result or its provenance
The firewall restricts who can write results; it does not certify that a method is right for your system. Choosing and judging a method remains scientific work.
Orbital-energy difference; not an excited-state calculation. Computed does not mean checked by a person: this result is stored as calculated and not reviewed.